This guide explains Drf Crefito, its clinical intent, and how professionals evaluate related treatments. Objectively, “DRF” and “CRE” are commonly used as shorthand across different medical and manufacturing contexts, so meanings depend on the exact field, documentation, and supplier specifications. Learn what to verify, what conditions to meet, and which quality checks support safe decision-making.
“Drf Crefito” may refer to different terminology depending on the specialty, country of use, and the supplier’s documentation. For that reason, the very critical action—before comparing options, discussing price, or planning a workflow—is to confirm exactly what the term denotes in your context (e.g., intended medical use, material category, regulatory status, and required handling conditions). This article provides an objective framework professionals use to evaluate such items and related procurement or clinical pathways, while emphasizing responsible verification over assumptions.
In many regulated or safety-critical environments, the “name” of an item is the least reliable part of the chain of evidence. Names can be abbreviated, translated, misspelled, or used inconsistently across hospitals, distributors, procurement catalogs, and even within the same organization. When “Drf Crefito” appears in a quotation, protocol draft, or internal checklist, the correct professional stance is to treat it as a pointer—something that must be mapped to unambiguous identifiers and supporting documentation—before decisions are made.
Beyond terminology, “Drf Crefito” may also be connected to a device category, an accessory bundle, a packaging configuration, a material grade, or a specific manufacturing variant. Each of those possibilities can carry very different handling requirements, shelf-life, inspection criteria, and compliance expectations. As a result, the “meaning” you validate should not stop at one definition; it should be validated across the practical dimensions that matter for safety, reproducibility, and audit readiness.
In healthcare, shorthand and abbreviated names can compress multiple concepts: the underlying device or material family, a manufacturing variant, a compatibility rule, and even a documentation set (labels, certificates, or test reports). When a term like Drf Crefito is encountered, industry experts typically treat it as a starting label—not as proof of performance. The safe approach is to obtain the “traceability packet” from the supplier: identification details, intended use statement, quality documentation, and any installation or handling requirements.
Additionally, variations in spelling and transliteration can occur across regions and vendors. An objective evaluation should therefore be based on verifiable identifiers (part numbers, batch/lot numbers, material specifications, and conformity documentation), rather than relying purely on the name “Drf Crefito.” In practice, it is not enough to ask, “What does it mean?” The better questions are:
Professionals also consider that terminology can be misused unintentionally. For example, a distributor might reuse a legacy abbreviation, a procurement team might copy and paste a shorthand entry from an older contract, or a clinician might reference a local label used in training materials. Even if the label is “mostly correct,” the “mostly correct” interpretation is the one that can fail during receipt inspection, audit review, or when a batch mismatch becomes important.
Because of this, evaluation should be evidence-based and structured. Experts typically build a mapping document internally that ties the label “Drf Crefito” to:
That mapping becomes the reference point for all subsequent steps: bid comparison, workflow planning, acceptance testing, and traceability logging.
Without assuming a single universal meaning, professionals commonly encounter Drf Crefito in contexts that involve one or more of the following categories:
Because different specialties may use different internal abbreviations, the exact interpretation of Drf Crefito must be grounded in your project’s technical brief or clinical protocol. That brief should ideally specify the patient population (where relevant), the clinical objective, constraints (time, setting, sterile/non-sterile requirements), and the operational environment. Without that context, the same shorthand label might be interpreted incorrectly.
To make this concrete, consider typical decision points where ambiguity causes downstream issues:
Professionals aim to prevent these issues by validating meaning early and continuously, not only at procurement time. “Drf Crefito” should trigger a structured verification workflow so that every later stage is supported by the same underlying evidence.
When teams compare product options associated with Drf Crefito, the comparison should start with documentation and end with the practical outcomes that matter to safety and reproducibility. Experts typically separate “cost” into multiple layers:
Even when pricing is attractive, teams often prioritize traceability and conformity evidence because these reduce operational uncertainty. “Cheaper” becomes “riskier” when it increases the probability of:
Where available, suppliers should provide supporting documents that align with applicable regulatory frameworks (e.g., medical device conformity expectations under relevant regional rules, as appropriate). The point of collecting documentation is not bureaucracy; it is to ensure that claims about intended use, performance, and safety can be substantiated and that your organization can repeat the same decision with the same evidence later.
Responsible evaluation also includes recognizing that different suppliers might use the same abbreviated label for different configurations. For example:
This is why you should compare options using structured evidence: product identity fields and document match criteria, not the shorthand name alone.
From an industry expert perspective, comparing suppliers for anything referred to as Drf Crefito should include a structured reliability review:
A key expert practice is to distinguish between “documentation available” and “documentation matched.” Documentation available might mean the supplier can provide general certificates on request. Documentation matched means those documents correspond to the exact SKU and batch/lot you will receive. For safety-critical decisions, matched documentation is the relevant standard.
Procurement reliability also includes responsiveness and clarity. A supplier who provides ambiguous answers, vague handling requirements, or inconsistent mapping between label and SKU is a reliability risk. Conversely, a supplier who can quickly provide a complete traceability packet with consistent identifiers is usually more reliable. Experts often summarize these reliability signals into practical procurement rules, such as:
These rules protect the organization from last-minute surprises.
Experts commonly use a checklist approach. For Drf Crefito related items, consider these objective checkpoints:
In real deployments, teams often discover additional “hidden variables” during these checkpoints. For example, a label might imply sterility, but the packaging certificate might indicate non-sterile supply requiring further processing. Or a supplier might list a broad intended use, but your protocol might require a narrower indication. Without verifying the intended use statement, compatibility errors can occur.
Another common issue is packaging and handling instructions that are specific to the product configuration. Even if the item identity is correct, a wrong storage condition—such as temperature range or exposure to moisture—can compromise quality. Therefore, the handling and storage checkpoint should be treated as essential, not as a training formality.
Finally, batch tracking should be considered early. It is not only a quality requirement; it is also a workflow requirement. Teams should ensure that their systems (electronic inventory systems, case carts, label printing, barcode scanning, and documentation templates) can capture the batch/lot identifiers provided by the supplier.
Healthcare quality systems place heavy emphasis on traceability because it enables investigation and corrective action if outcomes deviate from expectations. Globally, quality frameworks highlight the importance of identifying products, maintaining records, and supporting post-market or post-procedure review mechanisms (where applicable). While specific rules vary by jurisdiction and product category, the core principle is consistent: documentation and traceability reduce ambiguity in both day-to-day work and any later audit or review.
When traceability is robust, it becomes possible to answer key questions quickly:
When traceability is weak, those questions become difficult or impossible to answer. That can lead to unnecessary rework, clinical uncertainty, audit findings, or in worst cases, patient safety risk.
For medical device and health product governance, readers may consider guidance from recognized regulators and standards bodies, such as the U.S. FDA’s general principles on quality systems, or ISO-based quality management frameworks (for example, ISO 13485 in device contexts). These sources are widely used in industry to structure documentation and risk management practices.
Although this article does not replace jurisdiction-specific guidance, the professional mindset is consistent across many frameworks: establish controlled processes, ensure documented evidence for key steps, manage risk proactively, and maintain traceability from supplier receipt through to use. In that mindset, “Drf Crefito” is not a final category; it is an input label that must be mapped to the controlled product definition.
| Evaluation dimension | What to request/verify for “Drf Crefito” | Why it matters |
|---|---|---|
| Product identity | Exact item name, part number/SKU, lot/batch linkage, specification sheet | Prevents mix-ups between similar-labeled variants |
| Intended use | Supplier’s intended use statement and compatibility notes | Ensures the item fits your clinical objective |
| Documentation quality | Conformity/quality documentation appropriate to the item category | Supports audits and safety due diligence |
| Handling & storage | Temperature/humidity limits, shelf-life, preparation/opening steps | Reduces performance degradation risk |
| Quality management maturity | Supplier quality system overview; consistency across batches | Improves reproducibility and reduces uncertainty |
| Operational support | Training materials, technical contacts, and troubleshooting guidance | Improves correct usage in real workflows |
| Total cost view | Unit price plus documentation handling, storage requirements, and inspection time | Ensures “cheaper” doesn’t become “riskier” |
To make the table more actionable, teams often add two additional evaluation dimensions that connect directly to operational risk:
These extra dimensions help ensure that “Drf Crefito” remains stable and controlled over time, not just at one purchase event.
Below is a practical, requirements-first guide that procurement and clinical operations teams often use when a term such as Drf Crefito appears in technical notes, quotations, or case planning. Adjust the depth to your jurisdiction and product category.
Record where you saw “Drf Crefito” (internal protocol, vendor quotation, clinical plan) and any surrounding wording (device type, material, intended anatomical/site reference, or procedural context).
Professional tip: also capture the version or date of the document or quotation. If the meaning changes over time due to revisions, you will need to know which version drove the original interpretation.
Ask the supplier to map “Drf Crefito” to an official product identification set: part number/SKU, batch/lot labeling scheme, and the specification document(s).
This mapping should be in writing, not only verbal, so that it can be audited and used to resolve future discrepancies.
Compare the supplier’s intended use statement with your documented protocol. If there is any mismatch, clarify before procurement.
Also check whether the supplier specifies any limitations such as contraindications, operator requirements, patient suitability constraints, or environmental constraints (e.g., operating environment or procedure setting).
Assess whether the documentation matches the specific item and batch. Use a “document match” rule: the documents must correspond to what you will receive.
If the supplier provides generic documents, request the lot-specific traceability evidence. Where lot-specific documents are not available, document that limitation and decide whether acceptance is acceptable under your risk policy.
Verify storage conditions, shelf-life, packaging integrity expectations, and required preparation steps.
Confirm whether items require special storage (e.g., temperature control, desiccant control, humidity limits) and whether there are time limits after opening or reconstitution.
At delivery, verify identity, lot/batch numbers, and basic condition. Log the information immediately to support traceability.
Receipt inspection should include verifying that packaging labels align with the mapped SKU and that the item condition is consistent with handling requirements (e.g., packaging not compromised).
Make sure staff understand any procedure-specific steps associated with the product, including what to do if documentation or labeling is incomplete.
Training should be tied to the mapped product identity. If “Drf Crefito” is ambiguous in internal training materials, update the training documents so staff have a clear and consistent reference.
If any deviation occurs (e.g., handling condition differences), record it and follow your internal quality system steps.
Deviations should include what was observed, what was done to mitigate risk, and whether the item is released, quarantined, or rejected. The decision should be based on evidence, not assumptions.
To keep evaluations objective, teams should treat the following items as “non-optional” for anything referred to as Drf Crefito:
In practice, bypassing any of these can lead to a cascade of problems. For example, if documentation completeness is bypassed, acceptance might proceed without the evidence needed to confirm intended use. If compatibility verification is bypassed, workflow deviations may be tolerated informally until an incident triggers corrective action. If traceability is bypassed, incident investigations may become slow and incomplete. If handling compliance is bypassed, quality might degrade quietly without immediate visible change. And if audit readiness is bypassed, corrective actions may be delayed due to missing records or incomplete documentation.
Therefore, organizations typically define “stop points”—moments when procurement or clinical use cannot proceed until verification is completed. “Drf Crefito” should trigger those stop points because its meaning is not inherently self-evident.
Even though “Drf Crefito” may look like a single term, ambiguous mapping often manifests through a series of operational scenarios. Understanding those scenarios helps teams design better controls.
A supplier delivers items labeled under a shorthand like “Drf Crefito.” At receipt, your team confirms the packaging looks correct, but the lot/batch numbers presented on paperwork do not clearly match the batch labels. Alternatively, the supplier provides quality documents that do not clearly reference the specific batch. In that case, professionals usually do not accept the items into inventory for use. They quarantine the batch and request clarification or corrected documents.
This scenario is a common reason organizations build “document match” rules. It is also why traceability depends on both the physical labels and the documentation content.
A supplier describes “Drf Crefito” as suited for multiple procedure types. Your protocol, however, is narrower. The item might be technically usable outside the protocol, but the organization cannot assume that the intended use claim covers the exact clinical context or that local standards are satisfied.
Professionals respond by requiring an intended use statement review and compatibility confirmation. If the intended use does not match, they may choose a different product family or require an expert risk assessment and documented approval process, depending on local policy.
Staff might have used similar items before and assume the handling conditions are the same. But “Drf Crefito” might correspond to a variant with different shelf-life constraints, opening procedures, or packaging integrity expectations. If handling requirements are overlooked, quality can degrade before use.
To prevent this, teams require storage and handling verification as a formal checkpoint. Training should incorporate the correct steps and emphasize what must not be changed (e.g., storage temperature, exposure time, moisture protection).
Teams sometimes compare suppliers based on unit price quickly because the term “Drf Crefito” seems familiar. The comparison becomes incomplete because documentation quality, lot-specific evidence, and acceptance feasibility are not included.
Professionals avoid this by treating documentation and traceability readiness as part of the procurement scoring model. The procurement process should not proceed to final selection if the evidence is missing or ambiguous.
Even with a supplier mapping in hand, internal communication can be a weak link. To manage this, organizations often implement a controlled vocabulary approach for ambiguous terms. “Drf Crefito” should be replaced internally with mapped identifiers, or at least accompanied by a mapped definition.
A practical internal communication approach is to create a “Drf Crefito mapping record” that includes:
Then, in clinical protocols and procurement documents, staff can reference the mapped record rather than the ambiguous shorthand. This reduces confusion and prevents teams from repeating the same verification work each time “Drf Crefito” appears.
Another effective practice is to assign responsibility for interpretation. For example:
Clear ownership ensures that the mapping is not left to individual interpretation.
Risk management in healthcare is often about identifying uncertainty. When a term like “Drf Crefito” is ambiguous, uncertainty is built-in. Professionals handle this by converting uncertainty into a documented risk assessment step.
For example, a risk-trigger approach might involve:
This approach ensures the organization treats the ambiguous term as an operational risk trigger rather than as a harmless label.
When requesting clarification on “Drf Crefito,” you want questions that force a precise mapping and evidence delivery. Useful supplier questions include:
If the supplier cannot answer these questions with clear documentation, professionals generally consider the item “needs clarification” and delay final decisions until evidence is sufficient.
Receipt validation is where ambiguous terms are converted into controlled acceptance decisions. For “Drf Crefito,” a defensible acceptance process typically includes verifying identity, documentation match, and basic condition.
A structured receipt validation approach might include:
This creates a defensible chain of custody and ensures that “Drf Crefito” no longer exists as an ambiguous label once the item enters service.
Validation is not complete until it is reflected in standard work. Staff training and workflow documents should reference the mapped product identity rather than the ambiguous shorthand alone. This includes:
Training should be practical and scenario-based. For instance, staff should know what steps to take if “Drf Crefito” appears on a label but the part number does not match the internal mapping record. A clear escalation path prevents ad-hoc decisions under pressure.
One of the most overlooked aspects is that the meaning of a label can change over time. Even if “Drf Crefito” once mapped correctly, future shipments might correspond to a different SKU variant, manufacturing change, or packaging configuration change. Professionals handle this through document control and change management.
A robust change management process might include:
In controlled environments, change management is how you preserve the integrity of your evidence chain.
Answer: The exact meaning of “Drf Crefito” depends on the specialty and the supplier’s documentation. It may be an abbreviated term referring to a specific product family, variant, or workflow component. For accurate interpretation, request the official product mapping (part number/SKU, specification sheet, intended use statement) from the supplier or the source of the term.
Answer: Price alone is insufficient. Professionals compare total cost of quality, documentation completeness, handling requirements, and compatibility risk. A lower unit price can increase operational uncertainty if traceability or documentation is weaker.
Answer: Commonly requested materials include identity/specification sheets, batch/lot labeling information, and quality/conformity documentation appropriate to the product category. The goal is document matchability: the documents must correspond to the specific item and batch you receive.
Answer: Verify the intended use and compatibility notes against your protocol, and confirm whether the product requires specific systems, components, or procedural steps. Request technical support if any step in your workflow differs from what the supplier describes.
Answer: Typical reasons include mismatch between delivery labels and purchase documents, incomplete traceability information, ambiguous intended use, or missing/unclear handling instructions. Teams usually delay acceptance until documentation and specifications are verified.
Answer: Yes. Many medical or technical consumables have storage and handling constraints. The supplier’s storage requirements and shelf-life rules should be obtained before acceptance, and staff should follow opening and preparation instructions precisely.
Answer: Use guidance from recognized regulators and standards organizations relevant to your region and product category. For example, general quality system expectations are discussed by regulators such as the U.S. FDA, and quality system structures are commonly aligned with device quality standards such as ISO 13485 (where applicable). Always confirm the guidance that matches your jurisdiction and item classification.
In real deployments, the teams that move smoothly tend to run disciplined request-for-information (RFI) cycles. They do not treat “Drf Crefito” as a final technical definition. Instead, they require a mapping from the label to the spec sheet, then confirm that mapping supports safe use in their documented workflow. This approach also reduces rework—because it catches uncertainties early, before staff time, scheduling, and clinical steps become constrained.
Experts also recognize that “verification” is not a one-time action. Verification should occur at multiple points:
Another expert note is about how to avoid “interpretation by confidence.” A team might feel confident that they understand “Drf Crefito” because a colleague used it before. Professional practice requires evidence, not confidence. Even if the interpretation is correct, lack of evidence can still produce audit risk and operational uncertainty.
Where staff encounter ambiguous terminology repeatedly, experts often suggest building a “label resolution repository”—an internal database where ambiguous shorthand labels are mapped to controlled product records with dates and responsible owners. This repository helps prevent the same confusion from recurring across departments or shifts.
Whether you are evaluating Drf Crefito for clinical planning, procurement, or quality control, the professional path is consistent: confirm the exact meaning through official supplier documentation, compare options using documented specifications and traceability support, and enforce handling and record-keeping requirements. By focusing on verifiable evidence rather than naming ambiguity, teams can reduce operational risk and improve repeatability in practice.
Ultimately, the safest and most audit-defensible approach is to ensure that “Drf Crefito” is never treated as a complete technical definition. Instead, it should trigger the mapping and verification steps that convert a shorthand label into a controlled, traceable product identification that your organization can confidently use.
Striking the Perfect Balance: Navigating Premiums and Out-of-Pocket Expenses in Senior Insurance Plans
Explore the Tranquil Bliss of Idyllic Rural Retreats
How to Make Lasting Memories at Disneyland Attractions
Affordable Phones and Plans for Seniors
Affordable Full Mouth Dental Implants Near You
Unlock the Top Kept Secrets to Finding Your Ideal Dentist for Flawless Dental Implant Results!
Discovering Springdale Estates
Unveiling RS Sul Telecom Services
The Guide to Car Trading
Striking the Perfect Balance: Navigating Premiums and Out-of-Pocket Expenses in Senior Insurance Plans
Finding an optimal mix of insurance premiums and direct medical costs is crucial for effective healthcare budgeting. This discussion will delve into tactics to achieve a balance that aligns with your finances.
Unlock the Top Kept Secrets to Finding Your Ideal Dentist for Flawless Dental Implant Results!
Selecting an adept dentist is critical for ensuring the efficacy and your satisfaction during a dental implant procedure. This in-depth guide is designed to assist you in finding a dental expert who meets your personal requirements, guaranteeing that your path to dental implant success is seamless and fruitful.
Protect Your Perfect Smile: Master the Art of Defending Your Teeth Against Acid Attacks
Though they may tantalize the palate and invigorate the senses, foods and drinks with high acid content are a double-edged sword with regard to oral wellness. Continual exposure to these substances progressively strips away the protective enamel from our teeth, which can lead to increased sensitivity, changes in coloration, and a variety of other dental complications. This composition offers insight into relishing these zesty favorites while safeguarding your teeth from damage.
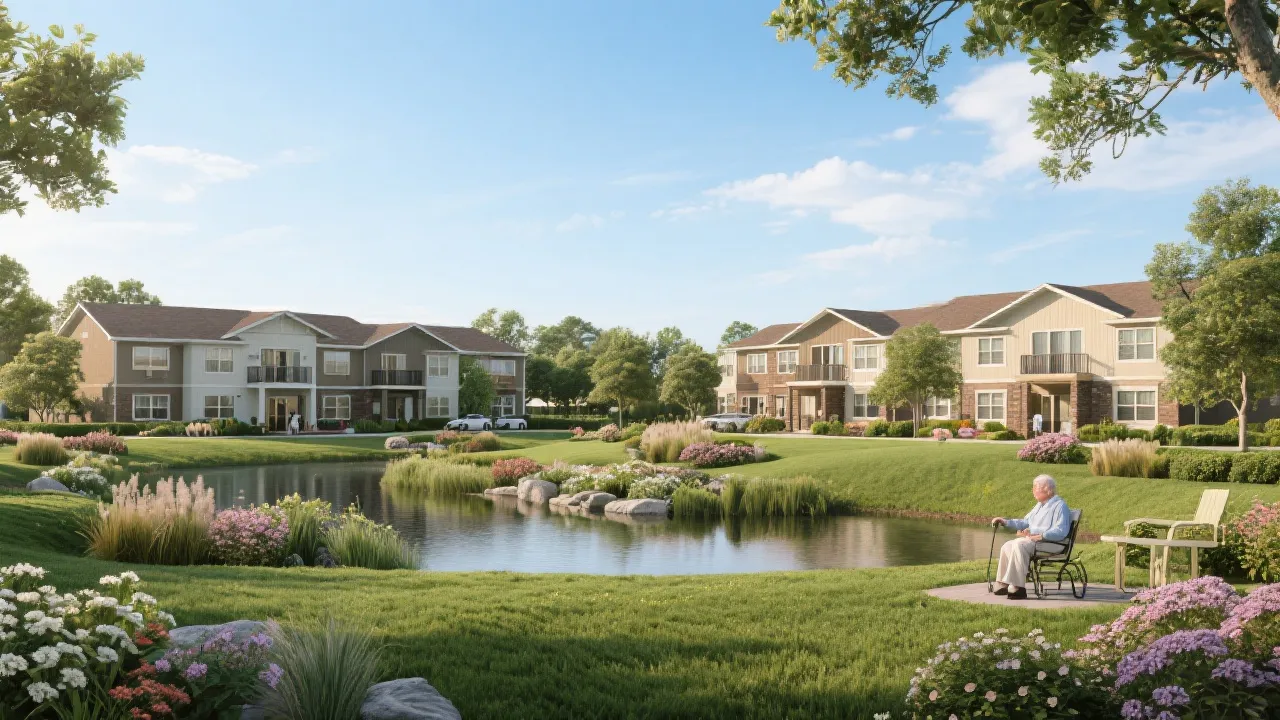
Discovering Heartis Eagle Mountain
Heartis Eagle Mountain is a prominent senior living community that offers a range of services and amenities aimed at enhancing the quality of life for its residents. Situated in a peaceful setting, it combines exceptional care with engaging activities, providing a holistic environment that caters to seniors' physical, emotional, and social needs.
Unveiling the Top Dental Implant Options for Seniors: Transform Your Smile with the Ultimate Guide to Restored Radiance
The significance of oral health cannot be understated for the elderly population as they are commonly afflicted with tooth loss due to a multitude of factors, including decay and gum disease. Dental implants have risen to prominence as a prime method of teeth replacement, acclaimed for their long-lasting nature, functional prowess, and striking similarity to our own teeth. This discussion delves deeply into the realm of dental implants to demonstrate why they are considered the optimal choice for aged individuals aiming to enhance their smiles and overall life quality.
Boost Your Defenses: 20 Proven Strategies for a Stronger Immune System
Having a robust immune system is essential to protect our bodies from harmful microorganisms and preserve our general health. Although some elements that affect our immunity are out of our hands, many lifestyle decisions can help us enhance our immune defenses naturally. This piece delves into 20 practical methods to empower your immune system, increasing your capacity to ward off diseases and sicknesses.